Resume template · Updated October 2026
Medical devices resume template
Short answer
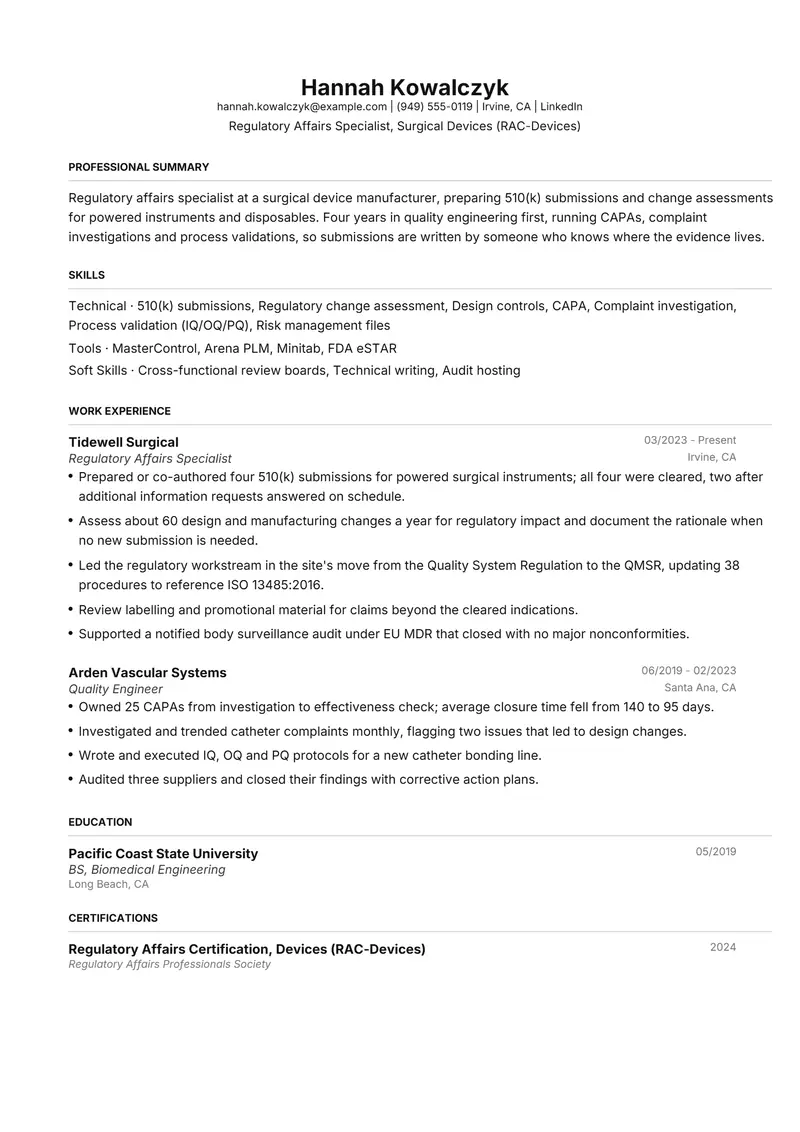
FAANGPath fits medical device roles, where exact regulations, standards and quality tools decide whether you are read further: its skills rows come before the jobs. The fictional sample is a regulatory affairs specialist at an Irvine surgical device maker who moved over from quality engineering, with 510(k) clearances, change assessments, the switch to FDA's QMSR, CAPA results and a RAC-Devices credential in her headline.
Download the medical devices sample
Hannah is a fictional regulatory affairs specialist. Replace every detail with your own before you send it.
The files are made in your browser by Resumity's own exporters, from this sample. The builder opens it as your own resume: click any line to replace it. Free plan, no watermark.
Why FAANGPath suits a medical devices resume
FAANGPath is the plain, centred layout meant for technical candidates, with labelled skill rows printed before work experience. Device hiring managers scan for exact terms (ISO 13485, design controls, CAPA, 510(k), process validation), and this order lets them find those terms before reading a single job.
Second choice: Meridian. Meridian shares the skills-first order with grey rules beside each heading, a slightly softer page for device sales or clinical specialist roles.
What goes in each section, in FAANGPath's order
FAANGPath prints its sections in a fixed order, the same in the PDF and the Word file. Here is that order for this sample, and what to put in each section.
- Professional Summary. Name the device type you work on (implants, diagnostics, capital equipment, disposables, software as a medical device), your function (R&D, quality, regulatory, manufacturing, clinical, sales) and the markets you cover. Device experience travels well, but reviewers check whether yours matches their risk class and regulators.
- Skills. Technical carries the quality and regulatory work you can do unsupervised: design controls, CAPA, complaint handling, risk management, process validation, submissions. Tools takes the eQMS, PLM and statistics software. Write standards with their numbers, since that is how they are searched.
- Work Experience. Device employers want evidence that your work held up under scrutiny: submissions cleared, audit findings closed, CAPAs that passed their effectiveness checks, validations approved, complaint trends reduced. Give counts and outcomes, and say when you hosted or supported an FDA inspection or a notified body audit.
- Education. Engineering and life science degrees dominate. Keep it to degree, field and school, adding a device design or validation capstone only in your first few years.
- Certifications. RAPS sets RAC eligibility at a bachelor's degree plus three years of regulatory or regulatory-related work, and counts quality assurance as related, which is why quality engineers often take RAC-Devices on the way into regulatory roles. FAANGPath prints it last, so repeat it in the headline.
The sample resume as text
The same content as the files above, so you can copy a line or compare it with your own.
Hannah Kowalczyk
Regulatory Affairs Specialist, Surgical Devices (RAC-Devices)
Irvine, CA | hannah.kowalczyk@example.com | (949) 555-0119 | linkedin.com/in/sample-hannah-kowalczyk
Regulatory affairs specialist at a surgical device manufacturer, preparing 510(k) submissions and change assessments for powered instruments and disposables. Four years in quality engineering first, running CAPAs, complaint investigations and process validations, so submissions are written by someone who knows where the evidence lives.
Experience
Regulatory Affairs Specialist, Tidewell Surgical, Irvine, CA (03/2023 to Present)
- Prepared or co-authored four 510(k) submissions for powered surgical instruments; all four were cleared, two after additional information requests answered on schedule.
- Assess about 60 design and manufacturing changes a year for regulatory impact and document the rationale when no new submission is needed.
- Led the regulatory workstream in the site's move from the Quality System Regulation to the QMSR, updating 38 procedures to reference ISO 13485:2016.
- Review labelling and promotional material for claims beyond the cleared indications.
- Supported a notified body surveillance audit under EU MDR that closed with no major nonconformities.
Quality Engineer, Arden Vascular Systems, Santa Ana, CA (06/2019 to 02/2023)
- Owned 25 CAPAs from investigation to effectiveness check; average closure time fell from 140 to 95 days.
- Investigated and trended catheter complaints monthly, flagging two issues that led to design changes.
- Wrote and executed IQ, OQ and PQ protocols for a new catheter bonding line.
- Audited three suppliers and closed their findings with corrective action plans.
Education
BS, Biomedical Engineering, Pacific Coast State University (05/2019)
Certifications
- Regulatory Affairs Certification, Devices (RAC-Devices), Regulatory Affairs Professionals Society (2024)
Skills
510(k) submissions, Regulatory change assessment, Design controls, CAPA, Complaint investigation, Process validation (IQ/OQ/PQ), Risk management files, MasterControl, Arena PLM, Minitab, FDA eSTAR, Cross-functional review boards, Technical writing, Audit hosting
How to make the sample yours
Bring your quality system language up to date
FDA's Quality Management System Regulation took effect on 2 February 2026, amending 21 CFR Part 820 and incorporating ISO 13485:2016 by reference. If you helped a site move from the old Quality System Regulation, say so; it is recent and specific.
Write submissions with outcomes
Under 21 CFR 807.81, a premarket notification goes to FDA at least 90 days before a device enters interstate commerce. Cleared 510(k)s, additional information requests answered and change assessments that avoided an unnecessary filing all belong in bullets, described by device type if the product is confidential.
Translate from pharma, automotive or aerospace
Validation, CAPA, supplier audits and change control exist in every regulated manufacturer. Map your terms onto device ones (batch records to device history records, deviations to nonconformances) and keep your numbers.
For field-facing roles
Clinical specialists and sales reps can trade submissions for cases supported, surgeons trained and territory growth, but should keep lines on adverse event reporting and promotional rules, which device employers expect everyone to respect.
Protect unreleased work
Unreleased products, inspection correspondence and audit findings are often confidential. Describe the device category, risk class and outcome rather than project names or document numbers.
Keywords to keep where they are true
Terms this kind of resume is usually searched for. Use the posting's exact wording, and only for work you have done.
- medical devices
- ISO 13485
- QMSR (21 CFR 820)
- design controls
- CAPA
- risk management
- 510(k)
- EU MDR
- complaint handling
- process validation
- change control
- internal audits
- technical documentation
- post-market surveillance
- RAC-Devices
Mistakes to avoid
- "Familiar with FDA regulations" without the part, submission or audit you actually worked on.
- Device code names, project numbers or inspection details your employer has not made public.
- Internal training presented as a certification next to real credentials such as the RAC.
- Standards you have only read about. Interviewers ask how you applied each one you list.
How the sample PDF reads in text extractors
We exported this sample with Resumity's PDF exporter in Chrome on 2026-10-29 and read the file back with three open-source text extractors. This shows whether software that turns a PDF into text can recover it; it is not a test of any commercial applicant tracking system. How we test templates.
| Extractor | Name, email, phone | Headings | Bullets intact |
|---|---|---|---|
| pdf.js 4.10.38 | All found | 5 of 5 | 9 of 9 |
| pdftotext (Xpdf) 4.00 | All found | 5 of 5 | 9 of 9 |
| pypdf 6.13.0 | All found | 5 of 5 | 9 of 9 |
Frequently asked questions
How do I get into medical devices from another industry?
Lead with what transfers: quality systems, validation, design control, supplier audits, cleanroom manufacturing, clinical knowledge. Learn the device vocabulary (ISO 13485, CAPA, 510(k)) and use it accurately; aerospace, automotive and pharma experience maps closely.
Should I list device classes on my resume?
Yes. Class II and Class III work differ a lot in regulatory effort, so "Class II powered instruments" or "Class III implants" tells a reader how close your experience is to theirs.
Is the RAC worth it for quality engineers?
It helps if you want to move into regulatory affairs, and RAPS counts quality assurance as regulatory-related experience. If you plan to stay in quality, a quality credential may fit your path better.
How do I describe confidential product work?
Use device category, risk class, market and outcome: "Class II powered surgical instrument, 510(k) cleared". Leave out code names, unreleased features and regulator correspondence.
Do device sales reps need regulatory keywords too?
Some. Sales and clinical specialist roles still involve adverse event reporting, cleared indications and promotional rules, so one line showing you work within them is worth keeping.
Will applicant tracking systems read this FAANGPath PDF?
The sample PDF read back with its contact details, all 5 section headings and all 9 bullets intact in three open-source text extractors (pdf.js, pdftotext and pypdf). Commercial systems such as Workday, Greenhouse or Taleo were not tested, and each employer configures its own.
Guides and templates for this resume
Sources, checked 29 October 2026
We opened every page below on that date. Pay, rules and requirements change, so check the source before you rely on a detail.
- U.S. Food and Drug Administration: Quality Management System Regulation (QMSR) (effective February 2, 2026; amends the device CGMP requirements of 21 CFR Part 820 and incorporates ISO 13485:2016 by reference)
- Legal Information Institute, Cornell Law School: 21 CFR 820.10 Requirements for a quality management system (manufacturers must document a quality management system that complies with ISO 13485 (incorporated by reference in 820.7 as ISO 13485:2016) and other applicable Part 820 requirements)
- Legal Information Institute, Cornell Law School: 21 CFR 807.81 When a premarket notification submission is required (premarket notification submitted at least 90 days before introduction or delivery for introduction into interstate commerce)
- Regulatory Affairs Professionals Society: Regulatory Affairs Certification (RAC) (RAC-Drugs and RAC-Devices; eligibility includes a bachelor's degree plus three years of regulatory or regulatory-related experience; quality assurance counts as regulatory-related)
More in resume templates by industry.